 Image 1 of 1
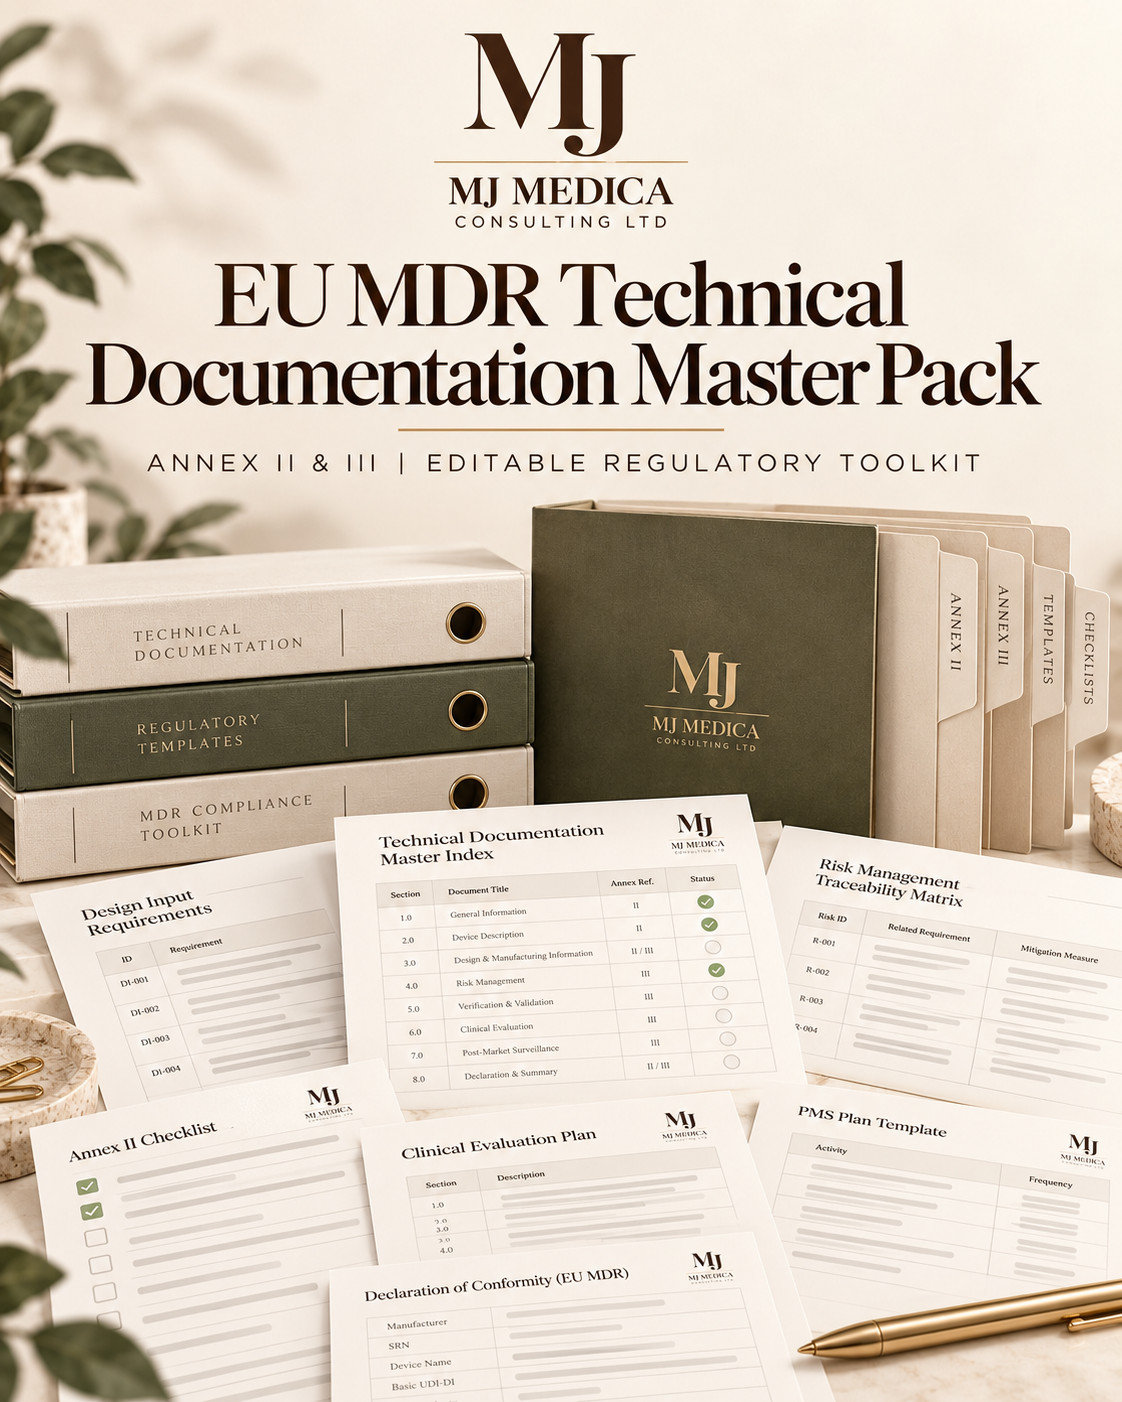
A comprehensive practitioner-developed EU MDR Technical Documentation toolkit designed to help medical device manufacturers structure, review, remediate and prepare Annex II and Annex III documentation for regulatory submission and ongoing lifecycle compliance.
The Master Pack brings together the core regulatory and quality documents needed to build a coherent, traceable Technical Documentation set rather than relying on disconnected templates.
The pack includes:
• Regulatory Strategy & Classification Assessment
• Technical Documentation Master Index
• Device Description & Specification template
• Intended Purpose & Claims Matrix
• MDR GSPR Compliance Matrix
• ISO 14971 Risk Management Plan and traceability tools
• Design & Manufacturing Evidence Index
• Verification & Validation Evidence Matrix
• Clinical Evaluation Plan / CER structure
• PMS Plan template
• PMCF Plan & Evaluation Report template
• PSUR / PMS reporting structure
• Labelling & IFU Review Checklist
• UDI / Registration Checklist
• Technical Documentation Gap Assessment
• Submission Readiness Checklist
• Master Remediation & Action Tracker
Designed to support:
• New MDR Technical Documentation development
• Legacy-device remediation
• Notified Body submission preparation
• Gap assessments and remediation programmes
• Internal regulatory reviews
• Change impact and lifecycle maintenance
• Cross-functional traceability between Regulatory, Quality, Risk, Clinical and Engineering evidence
Format: Editable Microsoft Word and Excel files supplied as a downloadable ZIP package.
Important: This toolkit is a structured regulatory starting point and must be adapted to the specific manufacturer, device, intended purpose, classification, conformity assessment route, target markets and applicable regulatory requirements. It does not by itself demonstrate conformity or replace device-specific regulatory judgement.
A comprehensive practitioner-developed EU MDR Technical Documentation toolkit designed to help medical device manufacturers structure, review, remediate and prepare Annex II and Annex III documentation for regulatory submission and ongoing lifecycle compliance.
The Master Pack brings together the core regulatory and quality documents needed to build a coherent, traceable Technical Documentation set rather than relying on disconnected templates.
The pack includes:
• Regulatory Strategy & Classification Assessment
• Technical Documentation Master Index
• Device Description & Specification template
• Intended Purpose & Claims Matrix
• MDR GSPR Compliance Matrix
• ISO 14971 Risk Management Plan and traceability tools
• Design & Manufacturing Evidence Index
• Verification & Validation Evidence Matrix
• Clinical Evaluation Plan / CER structure
• PMS Plan template
• PMCF Plan & Evaluation Report template
• PSUR / PMS reporting structure
• Labelling & IFU Review Checklist
• UDI / Registration Checklist
• Technical Documentation Gap Assessment
• Submission Readiness Checklist
• Master Remediation & Action Tracker
Designed to support:
• New MDR Technical Documentation development
• Legacy-device remediation
• Notified Body submission preparation
• Gap assessments and remediation programmes
• Internal regulatory reviews
• Change impact and lifecycle maintenance
• Cross-functional traceability between Regulatory, Quality, Risk, Clinical and Engineering evidence
Format: Editable Microsoft Word and Excel files supplied as a downloadable ZIP package.
Important: This toolkit is a structured regulatory starting point and must be adapted to the specific manufacturer, device, intended purpose, classification, conformity assessment route, target markets and applicable regulatory requirements. It does not by itself demonstrate conformity or replace device-specific regulatory judgement.